Contamination in pharmaceutical manufacturing occurs when unwanted microorganisms, chemical substances, or foreign particles enter a drug product during production or packaging. In tablet and capsule manufacturing, common sources include contaminated raw materials, airborne API dust, residues on shared equipment, guarnizioni usurate, lubrificanti, and inadequately cleaned product-contact surfaces. Cross-contamination involves material transfer between products, while packaging mix-ups represent a different quality risk.
Prevention requires controlling both contamination sources and transfer pathways. Facility segregation, HVAC airflow control, enclosed powder handling, localized dust extraction, hygienic equipment design, and validated cleaning provide complementary safeguards. Under GMP, these measures must reflect the product, manufacturing process, and assessed contamination risks.
What Are the Main Types of Contamination in Pharmaceutical Manufacturing?
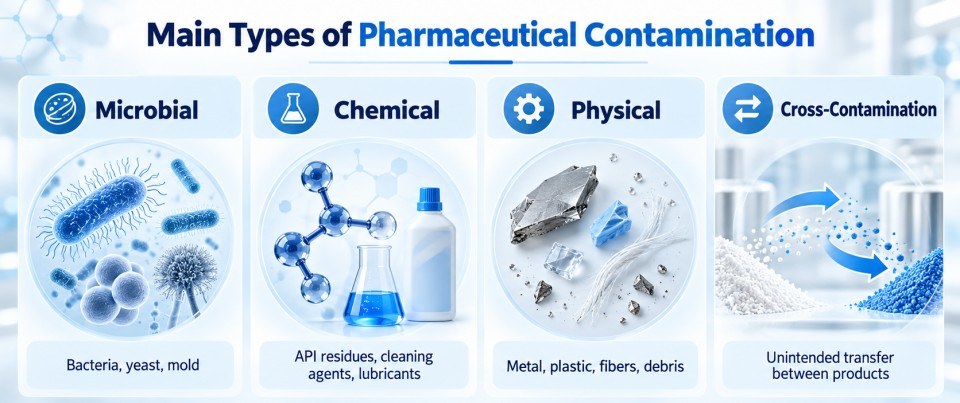
The three practical categories are microbial, chimico, and physical contamination, classified by the nature of the contaminant. Cross-contamination describes transfer from another material or product; confusione concern incorrect product or packaging identity. These risks can coexist but require different controls.
Microbial Contamination
Microbial contamination involves unwanted bacteria, yeasts, or molds introduced through raw materials, personale, acqua, attrezzatura, or the processing environment. Nonsterile tablets and hard capsules still have microbiological quality requirements appropriate to their formulation and intended use.
A cleaned powder hopper reassembled while wet illustrates the risk: retained moisture can support microbial survival or growth, particularly at poorly drained joints or seals. A suitable cleaning procedure must therefore address drying, storage before reuse, and inspection—not just visible powder removal. Moisture-sensitive formulations and hygroscopic excipients also require appropriate handling conditions.
Chemical Contamination
Chemical contaminants include residual active pharmaceutical ingredients (API), detergenti, lubrificanti, coolants, and unintended raw-material impurities. On shared equipment, residues can remain beneath a tablet press feed frame, in capsule dosing components, around hopper outlets, or behind worn gaskets.
If the next formulation contacts those residues, the result may be chemical cross-contamination. A lubricant leaking through a damaged seal presents a different route into the product zone. Neither problem necessarily produces visible debris. Controls therefore depend on cleanable geometry, integrità della tenuta, effective cleaning, and justified residue acceptance criteria.
Physical Contamination
Physical contamination introduces foreign particles such as metal wear debris, fractured punch or die components, gasket fragments, fibers, plastica, or packaging scraps. Likely points of entry include deteriorating machine parts, incorrectly assembled contact components, maintenance work, and exposed product transfer paths.
Ispezione dei componenti, manutenzione preventiva, suitable materials, and risk-appropriate foreign-particle detection address these sources. Pharmaceutical dust needs additional interpretation: a particle carrying an unintended API is also a chemical carryover hazard, even though it is physically a particle.
What Causes Contamination During Pharmaceutical Production?
The principal causes are contaminated incoming materials, powder release, retained residues on shared equipment, uncontrolled personnel or airflow routes, and mechanical wear or lubricant leakage. Risk increases when material escapes its intended path and remains available for later transfer.
Materie prime, Alimentazione, and Airborne API Dust
Incoming APIs and excipients require supplier qualification, identificazione, intact containers, suitable storage, and appropriate testing. These controls do not replace prevention during production.
Sampling, weighing, powder transfer, compressione della compressa, and capsule filling can generate airborne particles. Fine API-containing dust may escape an open hopper, a feeder-to-hopper joint, a tablet press feed frame, or a capsule dosing interface. It then settles on machine covers, utensili, gloves, or nearby surfaces, creating a residue reservoir that can be disturbed during cleaning, manutenzione, or the next batch.
Dimensione delle particelle, dustiness, flusso d'aria, and process speed affect release. Povero scorrevolezza della polvere can aggravate buildup but does not itself establish contamination.
Residues on Shared Equipment
Product-to-product carryover often begins in geometry that retains powder: tablet press feeders, pugni, muore, discharge chutes, capsule dosing discs, tamping-pin assemblies, hopper valves, transfer hoses, and gasket interfaces.
Concealed grooves and assembled joints can retain powder after visible surfaces appear clean. Vibration or startup movement can release it into the next batch. The inspection must cover product-contact and residue-retaining locations, not just the exterior.
Previous-product potency, toxicity, pulibilità, and the next product’s exposure determine the significance of carryover. A small residue from a highly potent API can pose a greater concern than a larger amount of a lower-risk substance.
Flusso d'aria, Personnel, and Mechanical Sources
HVAC supply air, local extraction, room pressure differentials, and personnel/material traffic determine where airborne powder travels. Neither positive nor negative pressure is correct for every operation: a pressure cascade must match the product-protection and containment objectives. Recycled air from dust-generating areas needs controls appropriate to the process.
Contaminated gloves, gowns, shared tools, and transferred machine parts can move residues even between rooms with otherwise suitable airflow. Nel frattempo, guarnizioni usurate, damaged gaskets, metal abrasion, cuscinetti, lubrificanti, and coolants create mechanical contamination sources. FDA 21 CFR 211.65 specifically addresses the need to keep operating substances from contacting drug products in a manner that affects quality.
| Source and transfer route | Typical tablet/capsule example | Control focus |
| Airborne API dust → surfaces → later batch | Powder escaping a feeder or dosing station | Source extraction, enclosure, pulizia |
| Residue → shared equipment → next product | Material retained beneath a feed frame or gasket | Disassembly access, validated cleaning |
| Moisture or microbes → product | Inadequately dried hopper or contaminated raw material | Asciugatura, igiene, material controls |
| Wear particles or lubricant → product zone | Damaged seal or worn mechanical component | Ispezione, manutenzione, mechanical isolation |
| Personnel/tools → another area | Dust transferred on gloves or tooling | Controlled flow, gowning, dedicated or cleaned tools |
How Can Contamination Be Prevented in Pharmaceutical Manufacturing?
Pharmaceutical contamination is prevented through facility segregation, suitable HVAC control, enclosed powder handling, localized dust extraction, effective equipment cleaning, personale addestrato, and risk-based verification. Each measure interrupts a different source or transfer pathway; none independently guarantees a contamination-free product.
Segregation, Flusso d'aria, and Material Routes
Segregate incompatible processes and establish controlled routes for incoming materials, sciupare, personale, and reusable equipment parts. Where justified, use dedicated rooms, airlocks, physical barriers, or separate handling systems to prevent cross-area transfer.
Air supply and exhaust positions must support containment rather than carry powder toward open products. For solid dosage lines, risk-based checks can include room pressure trends, extraction condition, housekeeping observations, and environmental sampling where justified. Do not automatically apply sterile-manufacturing environmental classifications to nonsterile oral solids.
Powder Containment and Source Extraction
Contain powder where it is generated. Closed transfers and sealed hopper connections reduce release during charging and conveying; enclosed processing zones limit escape around moving mechanisms; localized extraction captures dust near tablet feeders and capsule dosing stations before it disperses into the room.
Check extraction performance against the approved equipment setup, including airflow or differential-pressure indications where provided, blocked ducts, filter loading, raccolta delle polveri, and visible deposits. A dust collector does not by itself demonstrate high-containment performance for a potent API; such applications need a risk-appropriate containment strategy and demonstrated performance against defined requirements.
Pulizia, Cambio, and Line Clearance
Cleaning removes product and cleaning-agent residues. Cleaning validation demonstrates that the approved procedure consistently achieves justified acceptance criteria. The procedure must address difficult components, specified dismantling and reassembly, detergenti, rinsing if required, essiccazione, and protection before reuse.
Inspect representative hard-to-clean locations and use appropriate swab or rinse testing where required. A visually clean surface can still contain analytically detectable API residue. Nostro pharmaceutical cleaning validation guide explains acceptance limits, campionamento, and evidence in detail.
Line clearance is different: it verifies removal of previous products, etichette, documenti, and unsuitable packaging materials. The broader pharmaceutical production changeover process combines cleaning, sdoganamento, tooling adjustments, and restart checks; this article focuses on why those controls prevent contamination or mix-ups.
Personnel Controls and Ongoing Verification
Train operators to manage material spills, follow defined gowning and tool-cleaning rules, check machine condition, and document abnormalities. Verify seals after assembly, check residue-prone surfaces during clearance, and investigate recurring deposits rather than repeatedly cleaning without identifying their source.
Useful evidence includes cleaning records, residue results, extraction checks, maintenance findings, environmental monitoring where appropriate, and deviation trends. If a failure repeats, review the actual powder path, accesso per la pulizia, operator sequence, and equipment condition before changing an acceptance criterion.
How Does Pharmaceutical Equipment Design Help Reduce Contamination?
Equipment reduces contamination risk through sealed processing interfaces, source dust extraction, isolated lubrication, accessible product-contact parts, and geometry that minimizes retained residues. The design must match the specific powder route and be supported by inspection, manutenzione, and cleaning validation.
Riempimento di capsule: Sealing and Residue Access
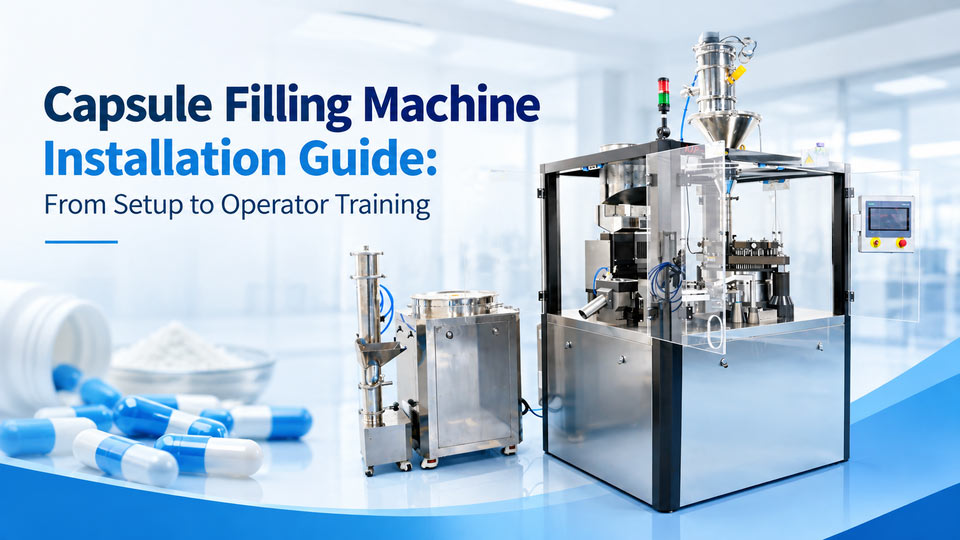
Nell'a macchina per il riempimento di capsule, powder travels from the hopper into dosing cavities and then the capsule body. The turntable, disco dosatore, perni di compressione, powder bed, and discharge components create interfaces where fines can accumulate. Powder that passes a turntable seal can enter internal mechanical spaces, where residue is more difficult to locate and remove during changeover.
Ruida Packing NJP-1500D automatic capsule filling machine is publicly described with turntable sealing and removable components for cleaning access. Its relevant pathway is dosing-zone dust → seal or joint → hidden residue → carryover risk at restart. Worn or misaligned seals can reopen that route.
Inspect seal contact areas, powder deposits around tooling, and removable contact components. Double seals do not mean zero dust or replace cleaning verification.
Pressa per compresse: Feed-Frame Dust and Lubrication Separation
UN comprimitrice rotativa repeatedly moves powder through the hopper, feed frame, die cavities, compression zone, and tablet discharge. Turret rotation, feeder movement, and tablet ejection can distribute fines around the die table and lower press area.
A compression enclosure limits escape; local suction captures dust near generation points. Inspect feeder edges, extraction inlets, die-table surfaces, and discharge guides for buildup. Filter loading or poor sealing can reduce capture even when the extractor runs.
Lubrication creates another route. Separate lubrication and product zones, inspect leaks, and replace worn seals.
Design igienico: Materiali, Dead Zones, and Disassembly
Cleanable geometry combines compatible contact surfaces, accessible joints, removable feeding parts, and fewer powder-retaining recesses. Critical areas include capsule dosing assemblies, tablet feeders, pugni, muore, and discharge paths.
Material grade alone is not a contamination control. 304, 316, and 316L stainless steels are not interchangeable specifications; suitability depends on actual machine documentation, corrosion exposure, formulazione, finish, and cleaning chemistry. Tool-free or modular disassembly may improve access and reduce changeover handling, but cannot establish cleaning effectiveness without a validated procedure.
| Equipment feature | Pathway it limits | What still needs verification |
| Turntable or dosing-area seals | Powder migration into mechanical cavities | Seal condition, adatto, hidden residue |
| Compression enclosure and local extraction | API dust escaping the press area | Flusso d'aria, filter state, powder deposits |
| Mechanical/lubrication isolation | Oil or grease contacting product | Leaks, sigilli, maintenance records |
| Removable contact parts and cleanable joints | Powder trapped in inaccessible recesses | Disassembly steps, campionamento, cleaning results |
| Packaging inspection and reject mechanisms | Wrong carton, missing leaflet or unit | Challenge testing and detection scope—not API carryover |
Conclusione
Contamination in pharmaceutical manufacturing is controlled by identifying what the contaminant is, where it originates, how it transfers, and which barrier stops it. In tablet, capsula, and packaging lines, powder containment, integrità della tenuta, cleanable tooling, flusso d'aria controllato, effective changeovers, and risk-based GMP verification address complementary risks. An enclosure or stainless-steel surface is only one part of the evidence; the real test is whether the complete process can repeatedly protect product quality.
Domande frequenti
What is contamination in pharmaceutical manufacturing?
It is the unintended presence or introduction of microbial, chimico, or physical contaminants in pharmaceutical materials, prodotti, attrezzatura, or processing environments. Risks arise during handling, manufacture, pulizia, e imballaggio.
What is the difference between contamination and cross-contamination?
Contamination describes unwanted material; cross-contamination specifically involves transfer from another material or product. Previous-product API residue entering a new formulation is an example of both chemical contamination and cross-contamination.
How can cross-contamination be prevented in tablet and capsule production?
Use risk-based segregation, trasferimento controllato della polvere, appropriate extraction, effective seals, cleanable product-contact components, validated cleaning, and changeover verification. The controls must fit the API, processo, e attrezzature.
How does pharmaceutical equipment design reduce contamination risk?
Seals restrict powder migration, enclosures and extraction capture dust, isolated lubricants reduce chemical entry, and accessible contact parts enable thorough cleaning. Performance still depends on correct setup, manutenzione, and verification.
Does GMP require dedicated equipment for every pharmaceutical product?
NO. Shared equipment may be acceptable when toxicological and process risk assessments support suitable controls. Dedicated equipment or facilities may be necessary when risks cannot be adequately controlled or demonstrated.
Riferimenti
- Commissione europea. Capitolo GMP dell'UE 3: Locali e Attrezzature E Capitolo 5: Produzione, particularly §§3.6, 3.14 and 5.18–5.22.
- NOI. FDA / eCFR. 21 CFR parte 211, particularly §§211.42, 211.46, 211.63–211.67 and 211.130.
- Commissione europea. Allegato GMP dell'UE 15: Qualificazione e convalida, Sezione 10.
- Organizzazione mondiale della sanità. Points to Consider When Including HBELs in Cleaning Validation.
- Consiglio internazionale per l'armonizzazione. Io Q9(R1): Gestione del rischio di qualità.